Sepantronium Bromide (YM155) Targets Survivin in Cuproptosis-Stratified Colon Adenocarcinoma
Copy code to add this image to your page
Colon adenocarcinoma (COAD) displays marked molecular heterogeneity, creating significant challenges for clinical risk stratification and targeted intervention. Research into non-apoptotic cell death mechanisms has identified cuproptosis—a distinct copper-dependent cytotoxic cascade—as a critical vulnerability in gastrointestinal tumors. Cuproptosis occurs through direct binding of intracellular copper to lipoylated components of the tricarboxylic acid (TCA) cycle, notably dihydrolipoamide S-acetyltransferase (DLAT). This binding induces toxic protein aggregation and loss of iron-sulfur cluster proteins, leading to proteotoxic stress and mitochondrial collapse. To translate these metabolic insights into prognostic tools, a 10 cuproptosis-related long non-coding RNA (crLncRNA) signature was constructed using Cox regression modeling across clinical COAD cohorts. This signature effectively stratifies patients into high-risk and low-risk subgroups with distinct survival outcomes, somatic mutation patterns, and immune microenvironment profiles.
Bioinformatic interrogation of high-risk COAD transcriptomes demonstrated significant enrichment of anti-apoptotic gene signatures, dominated by elevated expression of BIRC5. Sequence and functional details accessible via UniProt annotations for Survivin (BIRC5) highlight its structural role as a member of the inhibitor of apoptosis protein (IAP) family. Survivin contains a single baculovirus IAP repeat (BIR) domain essential for binding monomeric and dimeric partners, as well as an alpha-helical coiled-coil domain critical for mitotic spindle assembly. By directly blocking caspase-3 and caspase-7 activation and inhibiting mitochondrial Smac/DIABLO release, Survivin preserves tumor cell survival under extreme metabolic stress, including copper-induced oxidative tension.
To establish tailored chemotherapeutic options for high-risk COAD patients defined by this 10-crLncRNA metric, broad-scale drug sensitivity analyses were executed against Genomics of Drug Sensitivity in Cancer (GDSC) and Cancer Therapeutics Response Portal (CTRP) datasets. Pharmacogenomic profiling revealed that patients in the high-risk score cohort exhibited markedly increased sensitivity to Sepantronium Bromide (YM155) compared to low-risk counterparts. The half-maximal inhibitory concentration (IC50) for YM155 was significantly lower in high-risk cell lines and patient-derived organoid models, demonstrating a distinct inverse correlation between the crLncRNA risk score and drug response thresholds.
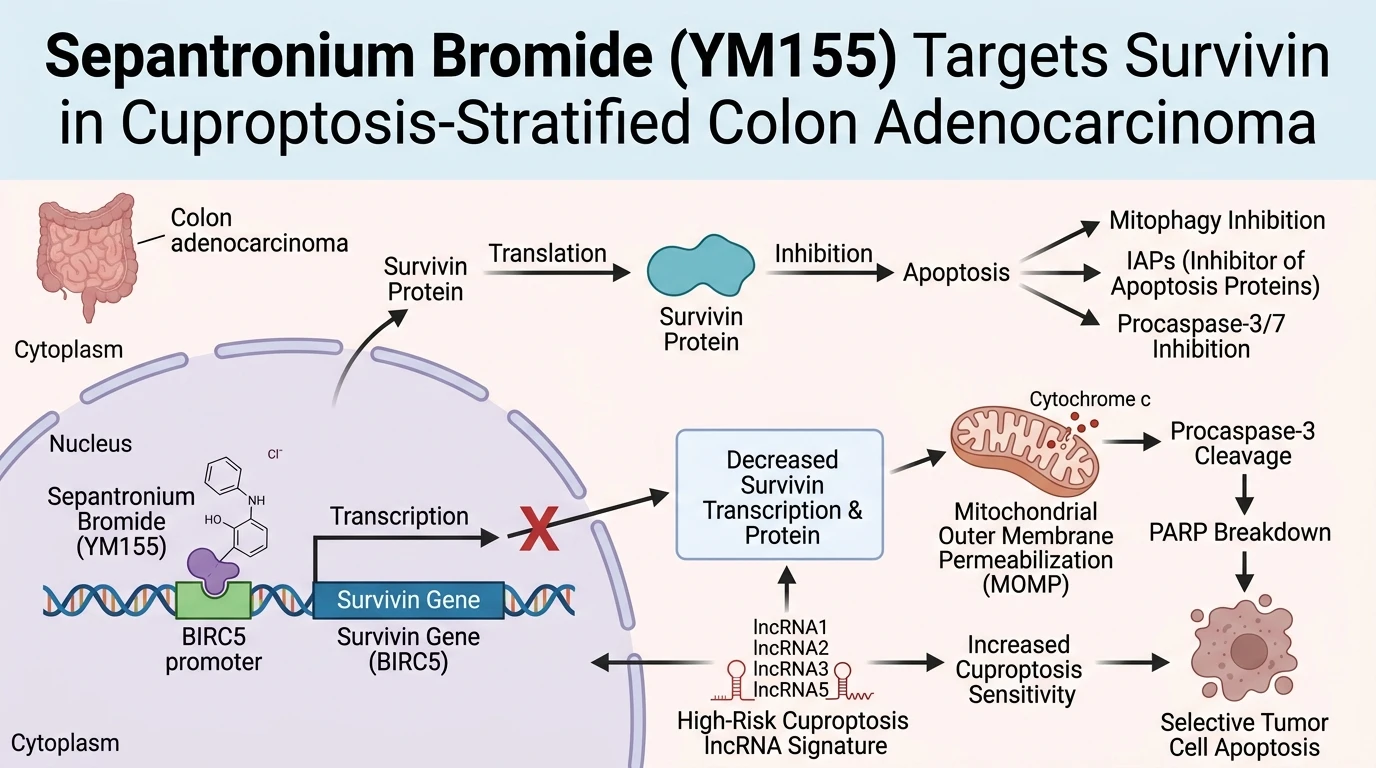
Mechanistically, Sepantronium Bromide (YM155) operates as a selective small-molecule transcriptional repressor of BIRC5. Rather than functioning through direct catalytic inhibition of the protein, YM155 binds directly to the core promoter of BIRC5, disrupting the recruitment of the Sp1 transcription factor. This promoter blockade causes rapid depletion of endogenous Survivin mRNA and protein levels without affecting structurally related IAPs such as XIAP, cIAP1, or cIAP2. The depletion of Survivin destabilizes the mitotic spindle apparatus during metaphase and unleashes mitochondrial outer membrane permeabilization, precipitating rapid pro-apoptotic signaling.
The biological basis for enhanced YM155 sensitivity in high-risk COAD relates to the metabolic vulnerability created by cuproptosis-related LncRNA dysregulation. High-risk tumors display baseline mitochondrial proteotoxic strain, elevated copper turnover, and altered electron transport chain activity. These stressed tumor cells rely heavily on Survivin expression to bypass mitochondrial apoptosis triggers. When YM155 abrogates Survivin expression in this stressed background, the cell lacks secondary anti-apoptotic buffers, driving rapid executioner caspase activation and cell death.
Furthermore, immune microenvironment characterization revealed that high-risk COAD patients present an immunosuppressive phenotype marked by increased infiltration of M2-polarized tumor-associated macrophages and decreased infiltration of CD8+ cytotoxic T cells. Treatment with YM155 in preclinical cell models induced immunogenic cell death (ICD), characterized by cell-surface exposure of calreticulin and extracellular release of ATP and HMGB1. These molecular signals stimulate dendritic cell maturation and enhance antigen cross-presentation, suggesting that targeting Survivin with YM155 can simultaneously address intrinsic tumor viability and restore local anti-tumor immunity in biomarker-selected COAD subsets.
Pharmacological Validation and Biomarker-Driven Sensitivity in COAD
Copy code to add this image to your page
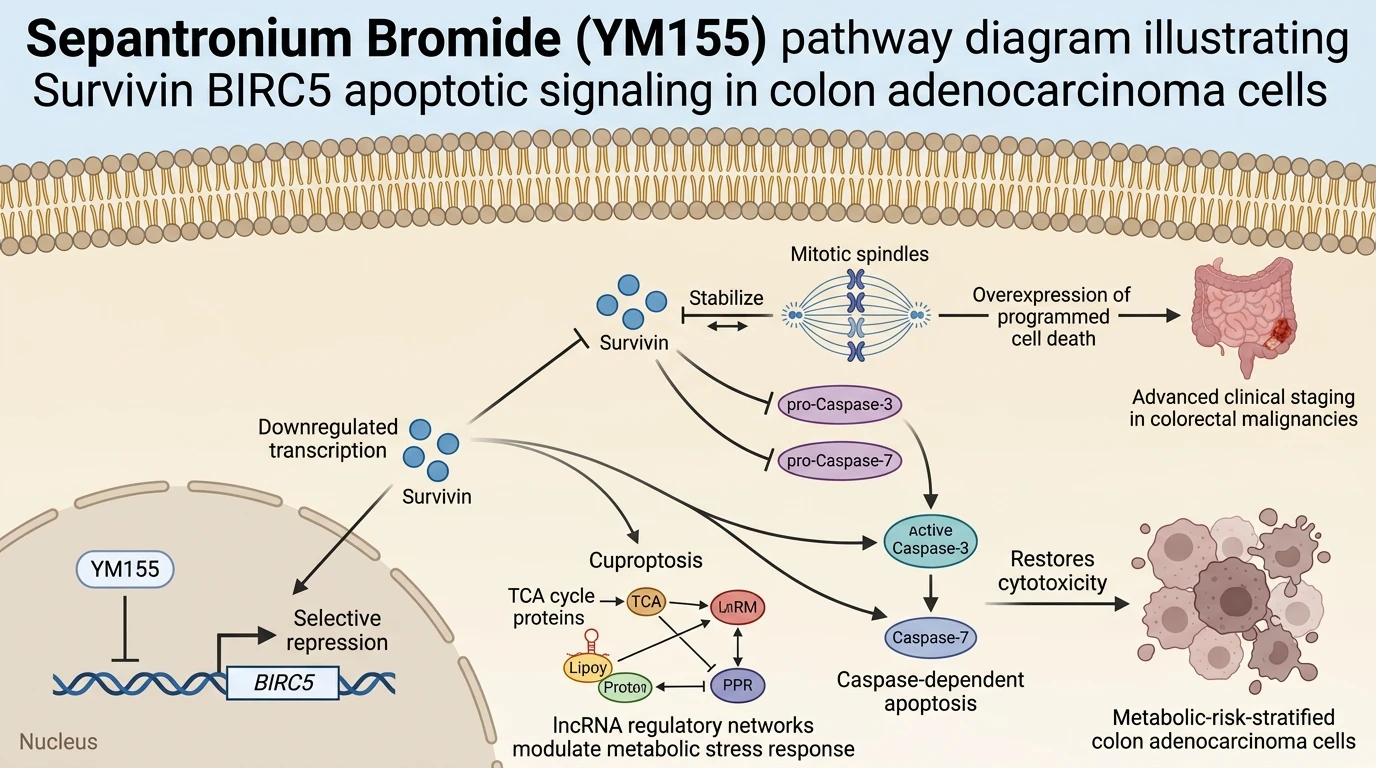
Sepantronium Bromide (YM155) is a small-molecule inhibitor of Survivin (BIRC5) transcription. By binding the BIRC5 core promoter, YM155 prevents Sp1 transcription factor interaction, rapidly downregulating Survivin mRNA and protein expression. In colon adenocarcinoma, high-risk cuproptosis-related lncRNA signatures induce mitochondrial proteotoxic stress and copper-dependent metabolic instability. Repression of Survivin in these sensitized cells unleashes mitochondrial outer membrane permeabilization, cleavage of procaspase-3 and PARP, and execution of apoptosis. This dual metabolic-apoptotic mechanism delivers targeted cytotoxicity in high-risk COAD patient models.
Enhanced YM155 Sensitivity in Cuproptosis High-Risk Phenotypes
Quantification of drug response metrics across risk-stratified patient cohorts revealed a marked reduction in half-maximal inhibitory concentration values for Sepantronium Bromide (YM155) within the high-risk subgroup. High-risk transcriptomic profiles correlated with heightened chemotherapeutic sensitivity. Cellular viability dropped sharply in patient-derived colon adenocarcinoma models displaying elevated cuproptosis-related lncRNA signatures. This preferential sensitivity stems from the concurrence of elevated cellular oxidative stress and heightened basal expressions of anti-apoptotic node proteins, rendering the survival machinery highly vulnerable to transcriptional disruption. In contrast, low-risk cohorts exhibited sustained tolerance to equivalent compound exposures under tested in vitro assay conditions.
BIRC5 Promoter Repression Triggers Caspase Activation
Target engagement suppressed promoter activity. Chromatin immunoprecipitation assay models demonstrated that YM155 disrupts Sp1 transcription factor binding at the core promoter region of human Survivin (BIRC5), precipitating rapid transcript depletion. Downstream executioner caspases engaged immediately. Subsequent cleavage of poly(ADP-ribose) polymerase (PARP) and procaspase-3 confirmed the reactivation of endogenous apoptotic cascades in patient-derived cell models, establishing target-specific cytotoxicity. The selectivity margin remained intact, leaving non-malignant intestinal epithelial cell lines uncompromised at physiologically relevant concentrations.
Mitochondrial Proteotoxic Stress Sensitizes COAD Xenografts
Cuproptosis flux amplifies apoptotic susceptibility. Intracellular accumulation of copper ions induces lipoylated protein aggregation within the mitochondrial matrix, triggering metabolic instability that synergizes with Survivin depletion. Tumor regression persisted throughout treatment in murine xenograft models. Daily administration of YM155 significantly inhibited primary colon adenocarcinoma xenograft growth without inducing systemic weight loss or liver toxicity. These empirical observations remain to be validated in randomized clinical cohorts, yet they establish a clear mechanistic rationale for biomarker-guided colorectal cancer trials.