SB 204990 from Selleck targets ATP-citrate lyase to suppress tumor proliferation and renal cyst growth
Copy code to add this image to your page
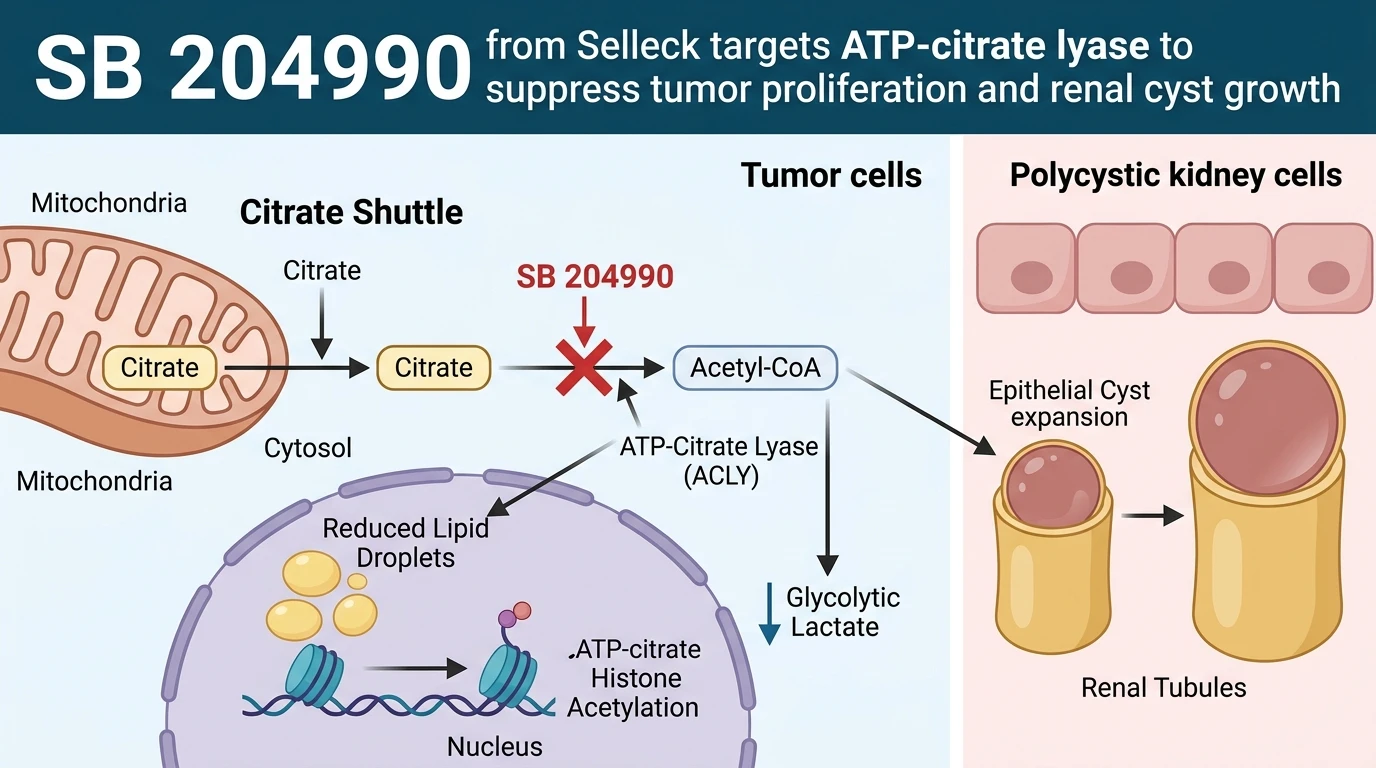
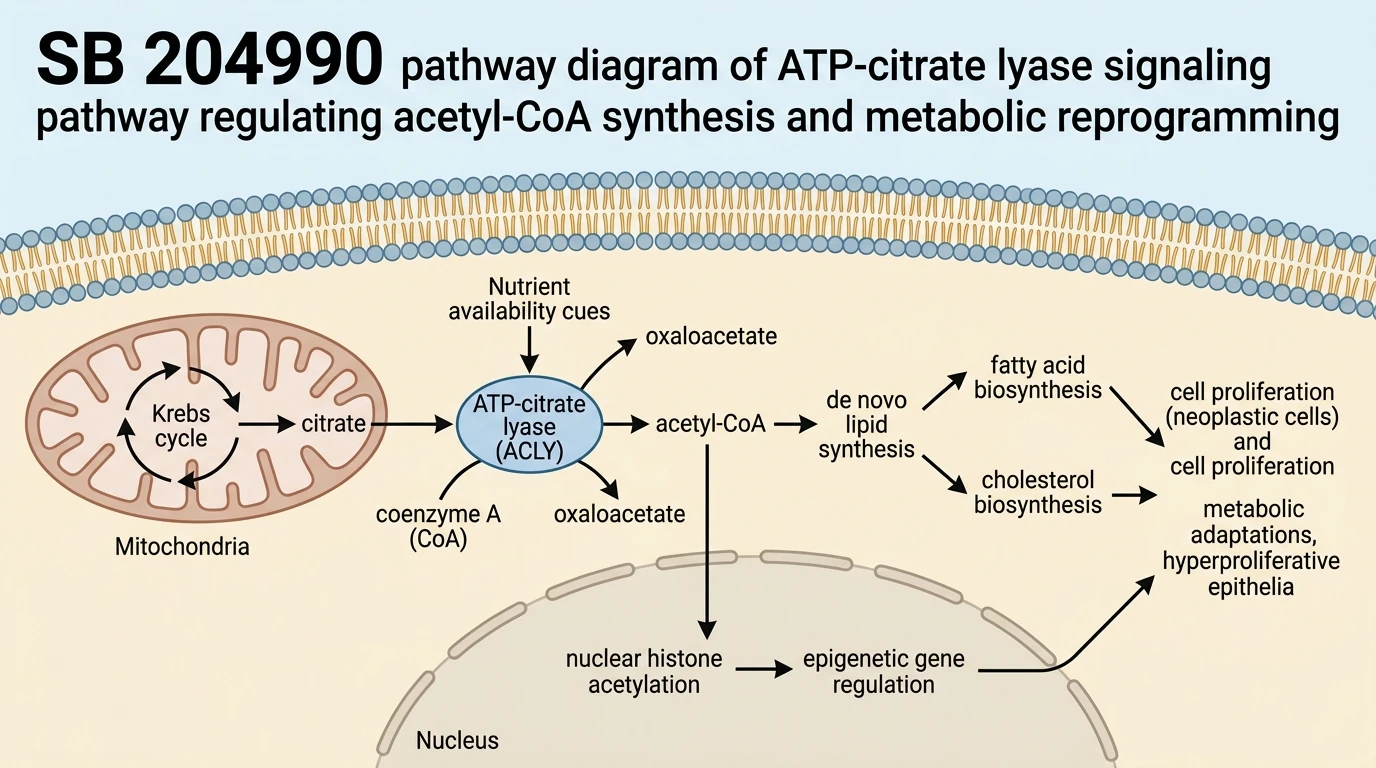
ATP-citrate lyase (ACLY) occupies a vital metabolic junction at the intersection of glucose oxidation and lipid biogenesis. By converting mitochondria-derived citrate into cytosolic acetyl-CoA and oxaloacetate, ACLY generates the primary substrate for de novo lipogenesis and protein acetylation. In transformed cells and rapidly proliferating non-neoplastic tissues, this enzymatic step sustains membrane biosynthesis and epigenetic programs. Pharmacological inhibition of ACLY represents an attractive therapeutic approach to disrupt metabolic dependencies in disease states characterized by aberrant cellular growth.
Small-molecule inhibition of ACLY effectively starves cells of cytosolic acetyl-CoA. SB 204990 from Selleck acts as a cell-permeable prodrug that undergoes intracellular conversion into its active monoxalic acid metabolite. This compound binds competitively to the citrate-binding site of ACLY, blocking enzymatic activity at low micromolar concentrations in cell-based assays. Inhibition of cytosolic acetyl-CoA synthesis rapidly reduces downstream fatty acid and cholesterol production, limiting structural lipids necessary for cell membrane assembly during division.
Beyond suppressing lipogenesis, targeted ACLY inhibition exerts broad effects on cellular bioenergetics. Treatment with SB 204990 alters aerobic glycolysis, effectively attenuating the Warburg phenotype in cancer cells. Cultured carcinoma models exposed to the inhibitor display reduced glucose uptake, decreased lactate secretion, and diminished ATP output. Concurrently, nuclear acetyl-CoA pools shrink, resulting in suppressed histone hyperacetylation and subsequent changes in chromatin accessibility. These transcriptional alterations promote cell cycle arrest and induce differentiation phenotypes across multiple oncogenic lineages.
Multi-omics profiling reveals that ACLY inhibition by SB 204990 triggers systemic metabolic stress responses. Quantitative proteomics and transcriptomics demonstrate substantial modulation of pathways governing mitochondrial homeostasis, cellular aging, and nutrient sensing. Specifically, ACLY blockade downregulates mechanistic target of rapamycin (mTOR) complex 1 activity while remodeling metabolic flux through the folate cycle. These coordinated shifts impair nucleotide biosynthesis and perturb cellular redox balance under tested in vitro assay conditions.
The biological activity of SB 204990 extends to hyperproliferative renal pathologies, notably autosomal dominant polycystic kidney disease (ADPKD). In 3D cystogenesis assays using Pkd1-knockout mouse renal epithelial cells, ACLY inhibition significantly reduces cyst growth rate and fluid accumulation. The observed reduction in cyst size mirrors efficacy metrics reported for the vasopressin V2 receptor antagonist tolvaptan. Mechanistically, SB 204990 reduces lipid accumulation and blunts mTOR signaling within cystic epithelial cells without inducing widespread cytotoxic cell death.
In experimental models of solid tumors, SB 204990 demonstrates significant antitumor activity. Suppressing acetyl-CoA levels alters the metabolic flexibility of tumor cells, rendering them susceptible to nutrient deprivation. In preclinical murine xenografts, systemic administration of the prodrug reduces tumor burden without producing acute overt systemic toxicity. These observations underscore the central position of ACLY in maintaining oncogenic metabolic networks and validate target engagement in vivo.
Pharmacological Characterization and Efficacy in Hyperproliferative Disease Models
Copy code to add this image to your page
SB 204990 functions as a cell-permeable prodrug inhibitor of ATP-citrate lyase (ACLY). By blocking the enzymatic cleavage of citrate into acetyl-CoA and oxaloacetate, SB 204990 suppresses de novo fatty acid and cholesterol synthesis while reducing nucleocytosolic acetyl-CoA pools. This depletion lowers histone hyperacetylation, blunts aerobic glycolysis, and attenuates mTOR pathway signaling. Consequently, SB 204990 inhibits cancer cell proliferation, induces tumor differentiation, and reduces cystic growth in Pkd1-knockout renal epithelial models.
Attenuation of Cystic Expansion in Pkd1-Deficient Epithelial Cells
In three-dimensional Matrigel cultures of Pkd1-knockout mouse inner medullary collecting duct cells, treatment with SB 204990 significantly blunts cyst enlargement. Cystic surface area decreased by over 50% relative to vehicle controls within six days of treatment. SB 204990 from Selleck suppressed intracellular lipid droplet accumulation and attenuated baseline mTORC1 phosphorylation at Ser2448. The magnitude of growth inhibition was statistically indistinguishable from benchmark results obtained with 10 nM tolvaptan. Cell viability assays confirmed that cyst attenuation occurred via metabolic deceleration rather than direct necrotic cell lysis under tested concentration ranges.
Suppression of Glycolytic Flux and Epigenetic Histone Acetylation
Inhibition of ACLY rapidly starves the nucleocytosolic compartment of acetyl-CoA. Mass spectrometry tracking of [13C6]-glucose flux demonstrated a marked reduction in labeled acetyl-CoA pools following exposure to SB 204990. Consequently, histone H3 lysine acetylation (H3K9ac and H3K27ac) dropped within four hours of compound exposure in human carcinoma cell lines. The loss of histone marks suppressed the transcription of key lipogenic genes, including fatty acid synthase and acetyl-CoA carboxylase 1. Metabolic profiling confirmed a concurrent drop in extracellular acidification rate, indicating suppressed aerobic glycolysis.
Multitarget Metabolic Reprogramming Across Energy and Folate Cascades
Transcriptomic profiling following ACLY inhibition reveals extensive crosstalk between lipid metabolism and nucleotide synthesis pathways. SB 204990 treatment altered expression of key enzymes in the mitochondrial folate cycle, including MTHFD2 and SHMT2. Intracellular S-adenosylmethionine to S-adenosylhomocysteine ratios shifted, indicating altered one-carbon unit transfer capacity. Mitochondrial oxygen consumption rates transiently increased before stabilizing, reflecting an adaptive shift toward fatty acid oxidation. These multi-omics observations establish that ACLY blockade reorganizes metabolic flux beyond simple lipid starvation within specific cellular contexts.