WY-14643 (Pirinixic Acid) Modulation of PPARα Signaling in Hepatic Lipid Metabolism
Copy code to add this image to your page
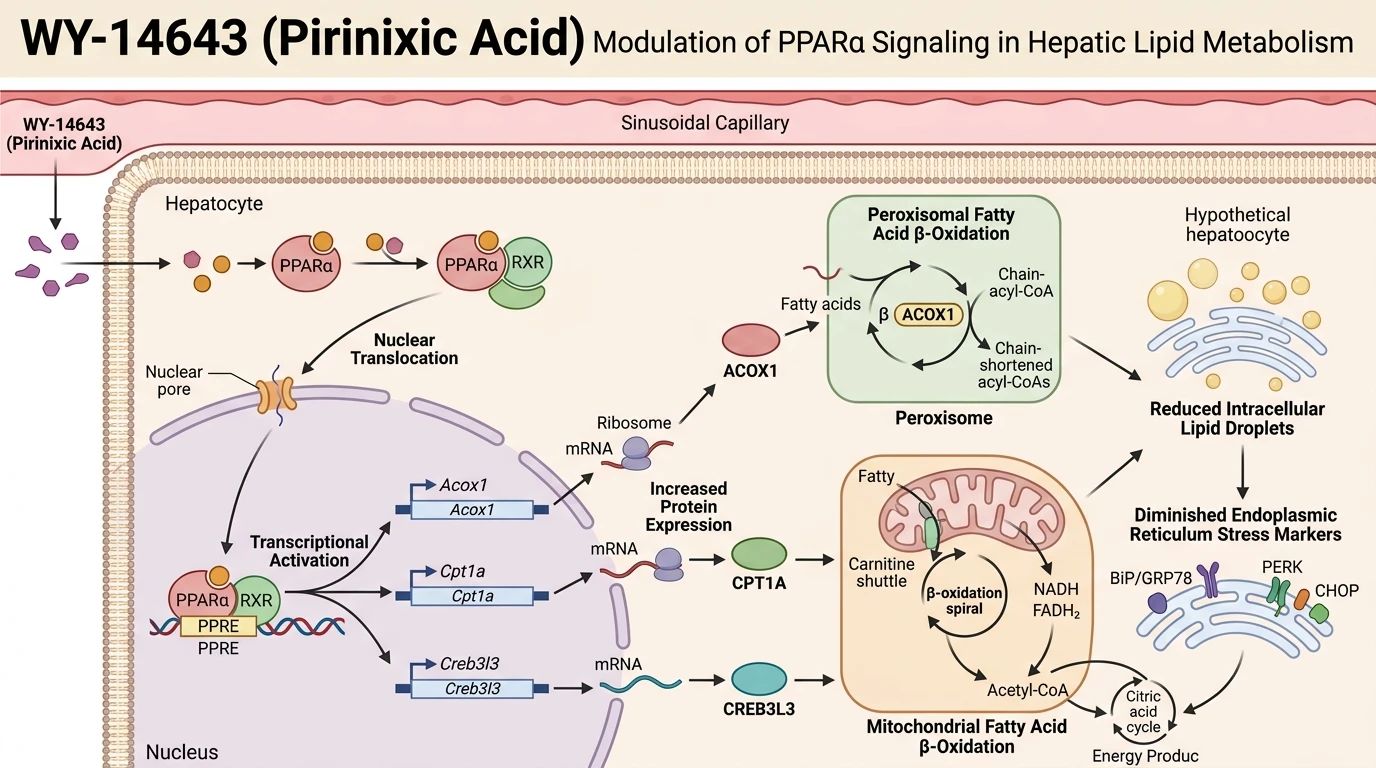
Peroxisome proliferator-activated receptor-alpha (PPARα) serves as a principal transcriptional driver of hepatic lipid catabolism and metabolic homeostasis. Classical full agonists, including fenofibrate and the prototypical synthetic ligand WY-14643 (Pirinixic Acid), have historically defined the mechanistic framework for PPARα-mediated transcriptional activation. These small molecules bind directly to the ligand-binding domain (LBD) of PPARα, prompting a specific conformational change that recruits nuclear receptor coactivators and promotes heterodimerization with retinoid X receptor alpha (RXRα). The resulting functional heterodimer binds peroxisome proliferator response elements (PPREs) located within promoter regions of target genes governing fatty acid uptake, peroxisomal β-oxidation, and mitochondrial transport.
Despite their broad utilization in biochemical studies, classical full PPARα agonists display variable transcriptional potency and off-target liability, driving interest in selective PPARα modulators (SPPARMα). A recent investigation evaluated the transactivation efficiency and lipid-lowering profile of K-877—a newly synthesized SPPARMα compound—in comparison to classical reference agonists, including Wy14643 and fenofibrate. In cell-based transactivation luciferase assays using human HepG2 hepatoma cells, K-877 exhibited exceptional transcriptional potency, registering an EC50 of 0.49 nmol/L. In contrast, classical agonists such as Wy14643 and fenofibrate required significantly higher concentrations to achieve equivalent transactivation levels in vitro.
The structural mechanics underlying SPPARMα activity differ fundamentally from conventional full agonism. Full agonists induce rigid stabilization of the activation function 2 (AF-2) helix, provoking robust coactivator recruitment and broad transcriptional induction across metabolic and non-metabolic gene networks. Conversely, SPPARMα ligands induce distinct receptor binding pocket geometries that favor selective cofactor interactions. In luciferase reporter systems, the WY-14643 PPAR interaction establishes high maximal efficacy, yet its relative transactivation potency remains orders of magnitude lower than that of SPPARMα molecules like K-877.
In vivo characterization in murine models further highlighted these distinct pharmacodynamic properties. Low-dose administration of K-877 (0.001% w/w in diet) induced targeted transcriptional upregulation of key hepatic fatty acid oxidation enzymes. Gene expression analysis revealed marked increases in acyl-CoA oxidase 1 (Acox1), which catalyzes the first rate-limiting step of peroxisomal fatty acid oxidation, and carnitine palmitoyltransferase 1A (Cpt1a), responsible for mitochondrial fatty acid transport. Additionally, transcript levels of cyclic AMP-responsive element-binding protein 3-like 3 (Creb3l3)—a key regulator of hepatic triglyceride secretion—were significantly elevated. These gene expression changes translated into rapid clearance of plasma triglycerides, surpassing the efficacy observed with standard fenofibrate diets.
Beyond circulating lipid control, PPARα activation provides structural protection against diet-induced hepatic pathologies. In mice fed a methionine-choline-deficient (MCD) diet, which induces non-alcoholic fatty liver disease (NAFLD) and steatohepatitis, treatment attenuated liver steatosis, inflammatory infiltration, and hepatocyte ballooning. Experimental evaluation in PPARα-null knockout mice demonstrated that these hepatoprotective effects are strictly target-dependent, as knockout cohorts exhibited no histological improvement. Furthermore, target engagement suppressed hepatic markers of endoplasmic reticulum (ER) stress, including phosphorylated eIF2α and CHOP expression, thereby stabilizing cellular protein homeostasis under metabolic strain.
Comparative molecular profiling demonstrates how refined ligand-receptor interactions shape metabolic responses in hepatic tissue. While reference tools such as WY-14643 (Pirinixic Acid) remain indispensable for basic characterization of receptor dynamics, high-potency selective modulators illustrate how subtle conformational variations yield distinct biological outcomes. Insights gained from nuclear receptor research initiatives continue to clarify how differential cofactor recruitment influences gene network architecture and metabolic resilience. Consequently, comparative studies between classical agonists and novel SPPARMα molecules provide essential benchmarks for dissecting the molecular regulation of hepatic lipid catabolism and cellular stress pathways.
Comparative Pharmacodynamics and Hepatic Gene Regulation
Copy code to add this image to your page
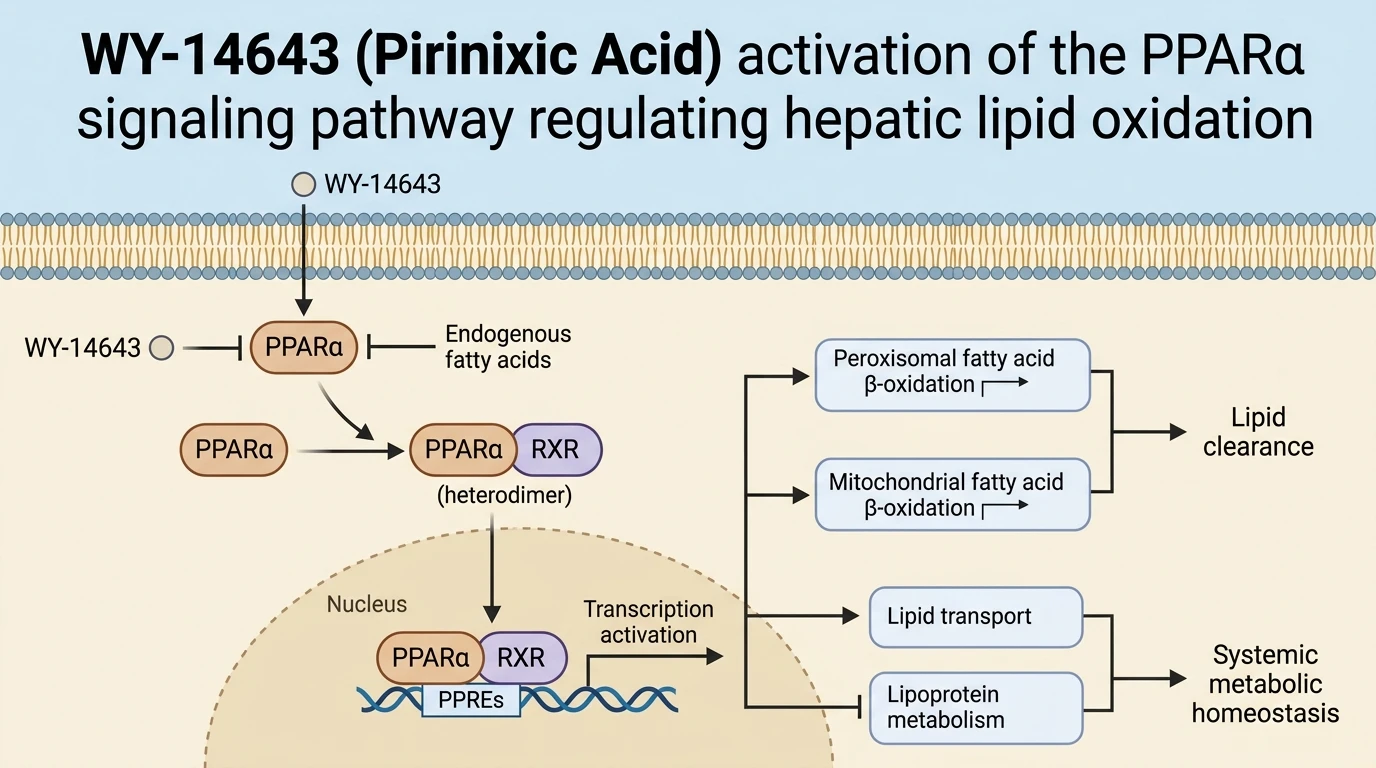
WY-14643 (Pirinixic Acid) and selective modulators act by engaging the peroxisome proliferator-activated receptor-alpha (PPARα) ligand-binding domain. Receptor activation drives heterodimerization with RXRα and binding to peroxisome proliferator response elements (PPREs). This transcriptional complex upregulates crucial hepatic fatty acid oxidation genes, including Acox1 and Cpt1a, alongside metabolic regulators such as Creb3l3. Enhanced β-oxidation accelerates triglyceride clearance, while simultaneously alleviating endoplasmic reticulum stress and hepatic steatosis in experimental non-alcoholic fatty liver disease models.
Luciferase Transactivation Kinetics in HepG2 Hepatocytes
In cell-based transactivation reporter assays using human HepG2 cells, K-877 demonstrated exceptionally high potency with an EC50 of 0.49 nmol/L. Classical agonists required micromolar concentrations. The WY-14643 PPAR interaction generated robust maximal transactivation, serving as an effective benchmark for full ligand efficacy. Coactivator recruitment patterns account for these observed potency disparities. High-affinity receptor engagement stabilizes specific binding pocket conformations, driving enhanced reporter gene transcription at sub-nanomolar concentrations.
Upregulation of Hepatic Fatty Acid Oxidation Transcripts in Vivo
Murine dietary intervention assays revealed significant induction of metabolic target genes following low-dose SPPARMα treatment. Dietary administration of 0.001% K-877 increased hepatic mRNA expression of Acox1 and Cpt1a. Transcript levels rose sharply. Expression of Creb3l3—a master regulator of hepatic triglyceride metabolism—elevated in parallel with peroxisomal beta-oxidation markers. These transcriptional shifts directly induced rapid plasma triglyceride clearance. Standard fenofibrate regimens required higher relative doses to match these quantitative shifts.
Attenuation of MCD-Induced Hepatic Steatosis and ER Stress
Evaluation in methionine-choline-deficient diet models established the therapeutic boundary of targeted PPARα modulation. Treatment attenuated hepatic triglyceride accumulation and tissue inflammation in wild-type mice. Protective efficacy vanished in PPARα-knockout cohorts. This loss of activity confirms absolute target dependence under tested dietary conditions. Furthermore, selective modulation suppressed hepatic markers of endoplasmic reticulum stress, including phosphorylated eIF2α pathways. Selective modulation achieved robust tissue protection.