SMS121 from Selleck blocks CD36-mediated fatty acid transport in acute myeloid leukemia cells
Copy code to add this image to your page
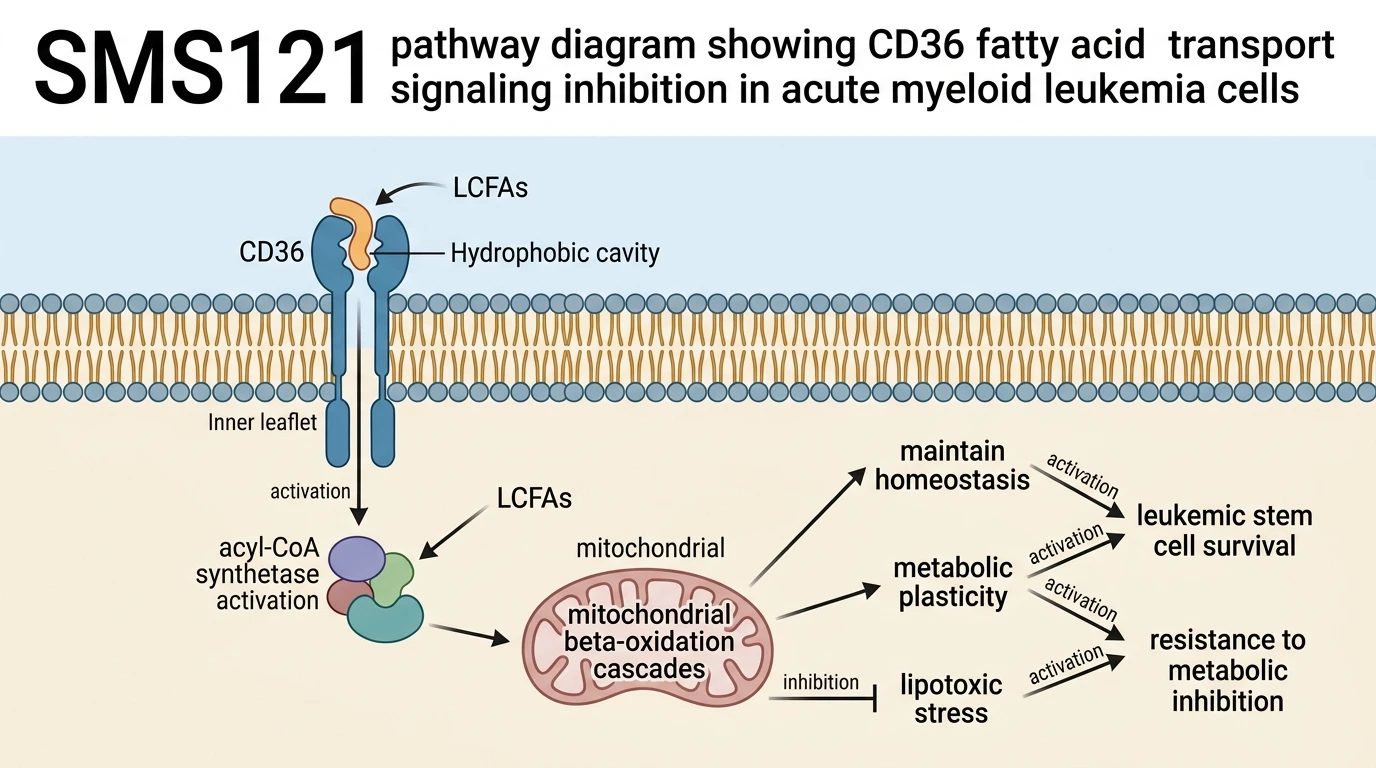
Targeting cancer cell metabolism has emerged as a compelling paradigm in modern oncology, particularly within hematologic malignancies where metabolic flexibility supports drug resistance and survival. Acute myeloid leukemia (AML) cells, especially therapy-resistant leukemia stem cell sub-populations, often exhibit a marked reliance on exogenous lipid utilization. The transmembrane glycoprotein CD36, functioning as a primary fatty acid translocase, plays an indispensable role in importing long-chain fatty acids across the plasma membrane into the cytoplasm. Elevated surface expression of CD36 correlates with adverse outcomes, enhanced metabolic adaptation, and chemotherapeutic resistance in AML patients. Consequently, pharmacological attenuation of CD36 translocase activity represents an attractive therapeutic strategy to selectively starve leukemic blasts of essential lipid substrates.
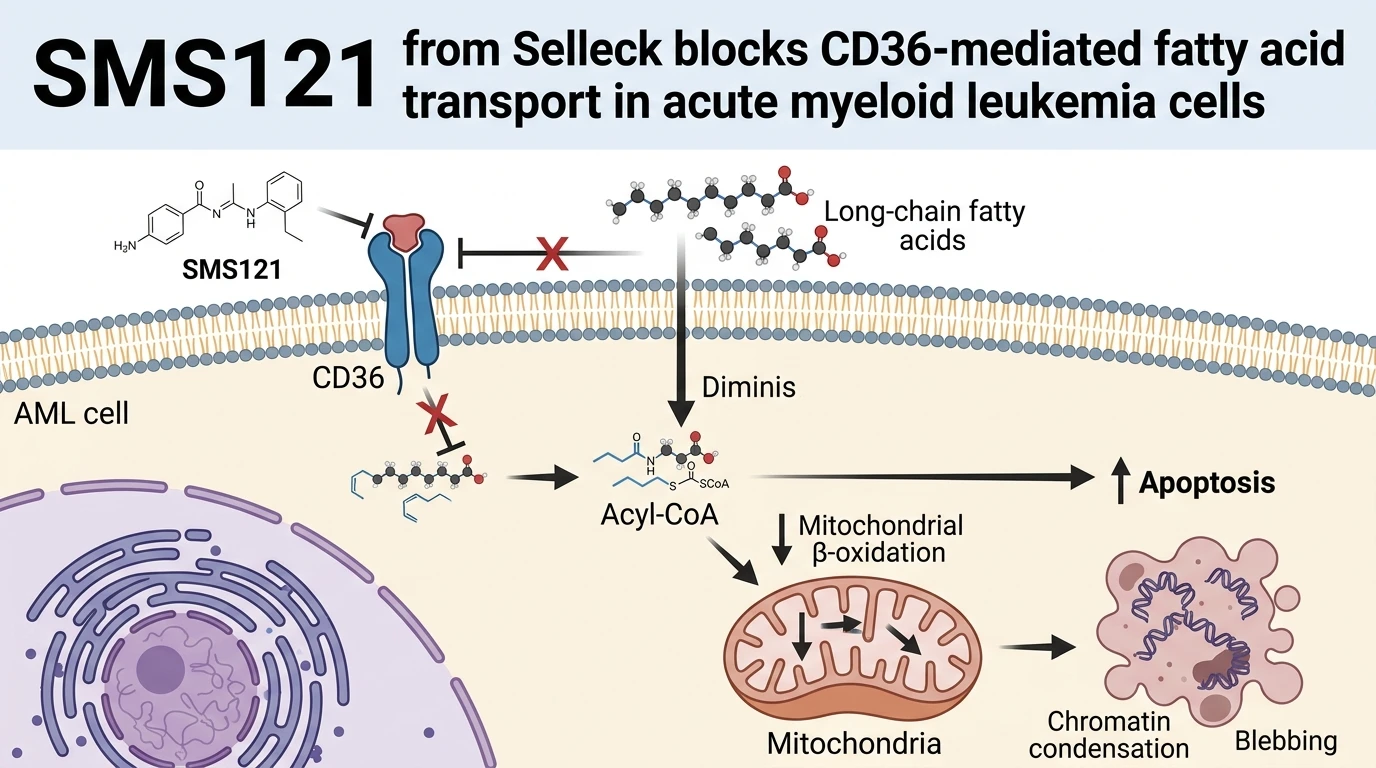
SMS121 from Selleck represents a novel, drug-like small-molecule inhibitor specifically identified to disrupt CD36-mediated lipid transport. Discovered through receptor-based consensus virtual screening coupled with extensive molecular dynamics simulations, SMS121 was optimized for high-affinity engagement within the structural architecture of the target protein. Virtual screening libraries were evaluated against the extracellular domain of CD36, applying consensus scoring functions to filter for optimal steric fit, hydrogen bonding potential, and hydrophobic interactions. Subsequent nanosecond-scale molecular dynamics trajectories demonstrated that SMS121 stably embeds within the central hydrophobic binding cavity of CD36. This precise engagement physically occludes the channel through which long-chain fatty acids traverse the outer cellular envelope. Biophysical characterization utilizing surface plasmon resonance and microscale thermophoresis established that SMS121 binds directly to recombinant human CD36 with low micromolar affinity, exhibiting an equilibrium dissociation constant (KD) of approximately 5 µM.
To evaluate the functional consequences of CD36 blockade in cellular models, investigations focused on acute myeloid leukemia cell lines displaying elevated CD36 surface density, such as the KG-1 lineage. In these high-expressing AML models, treatment with SMS121 resulted in a robust, dose-dependent reduction of cellular long-chain fatty acid uptake, yielding an IC50 value of 164 µM under standardized assay conditions. Radiolabeled and fluorescent fatty acid tracking assays confirmed that SMS121 selectively restricts the internalization of substrates such as palmitate and oleate. Inhibition of fatty acid uptake subsequently deprived the leukemic cells of critical substrates necessary for mitochondrial beta-oxidation, carnitine palmitoyltransferase-1 (CPT1) dependent ATP generation, membrane phospholipid synthesis, and lipid-derived signaling cascades.
The downstream functional impact of this metabolic blockade was evidenced by a parallel, dose-dependent loss of AML cell viability and proliferative capacity, with a measured IC50 of 156 µM. Notably, the close numerical correspondence between the IC50 for fatty acid transport inhibition (164 µM) and the IC50 for cytotoxicity (156 µM) indicates that the primary mechanism driving cell death is directly coupled to the suppression of lipid import rather than off-target non-specific toxicity.
To rigorously confirm target specificity and rule out non-specific cytotoxic effects, competitive rescue experiments were conducted in vitro. Addition of exogenous oleic acid or bovine serum albumin to the culture medium effectively neutralized the cytotoxic impact of SMS121, fully restoring AML cell viability and proliferation metrics. This complete phenotypic rescue demonstrates that SMS121-induced cell death is driven specifically by lipid starvation resulting from CD36 inhibition. When excess fatty acids or lipid carriers are introduced, the metabolic deficit is bypassed, confirming that the compound acts as a selective metabolic inhibitor. In summary, the characterization of SMS121 validates the feasibility of targeting CD36-mediated lipid uptake with small-molecule therapeutics, establishing a foundation for targeted metabolic therapy in acute myeloid leukemia.
Therapeutic Evaluation and Pharmacological Characterization
Copy code to add this image to your page
SMS121 is a small-molecule inhibitor targeting the lipid translocase CD36. By binding to the central cavity of CD36 with a KD of ~5 µM, SMS121 inhibits long-chain fatty acid uptake (IC50 = 164 µM) in CD36-expressing acute myeloid leukemia cells. This block in fatty acid transport impairs cellular lipid metabolism, reducing leukemic cell viability and proliferation (IC50 = 156 µM). Exogenous lipid supplementation rescues this phenotype, confirming a specific metabolic mechanism.
Virtual Screening Identifies Low-Micromolar CD36 Cavity Inhibitor
Receptor-based consensus virtual screening identified SMS121 as a drug-like antagonist of lipid translocase CD36. Dynamic simulations confirmed stable engagement within the central binding cavity. The molecule exhibited a dissociation constant (KD) of 5 µM in biophysical assays. Small-molecule inhibition directly disrupts fatty acid docking at the plasma membrane.
Fatty Acid Transport Blockade Induces Selective Cytotoxicity
In high-CD36 KG-1 leukemia cells, long-chain fatty acid uptake dropped significantly following exposure. The measured IC50 reached 164 µM under tested in vitro conditions. Cellular proliferation halted in tandem with an IC50 of 156 µM. Exogenous oleic acid or bovine serum albumin supplementation completely rescued cell viability. SMS121 from Selleck confirms that CD36 inhibition exposes a critical metabolic vulnerability in acute myeloid leukemia.